Ensembl opens nine billion predictions to labs
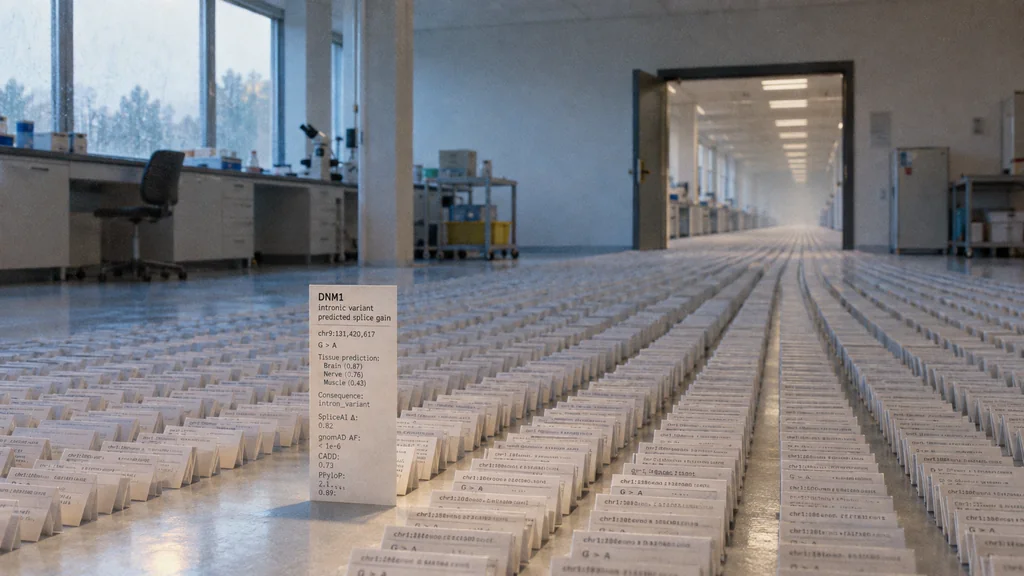
One predicted mutation rises, but only an experiment can confirm it.
Image composition · tobriefTake the human reference genome, a standard copy of our roughly three billion DNA positions. At each position sits one of four chemical letters. Swap any letter for one of the three alternatives, and you get a hypothetical mutation. Google DeepMind's AlphaGenome model did exactly that, running predictions on all possible single-letter substitutions to produce about nine billion entries, each carrying an average of roughly 27,000 molecular predictions across tissues and cell types (DeepMind). The resulting Atlas also covers more than 100 million short insertions and deletions already catalogued in population databases (Atlas preprint).
None of these entries are experimental measurements. They are what the model predicts would change if each swap occurred. The question is whether those predictions help researchers find variants that matter.
A buried variant, a brain-specific clue
The most concrete test case involves a person with severe epileptic encephalopathy whose genetic sequencing had not yielded an answer. Researchers at the Broad Institute, working with the GREGoR Consortium and the AlphaGenome team, used the Atlas's summary score, called AVI (AlphaGenome Variant Impact), to reprioritise the patient's variants. It elevated a change buried deep inside an intron of the gene DNM1 (Fortune). An intron is a stretch of DNA that gets cut out before a gene's instructions are translated into protein. Analysts rarely look there, and previous RNA analysis from blood had been inconclusive because DNM1 is barely active in blood cells.
A separate attribution analysis estimated that 69% of the AVI signal came from predicted splicing effects, the process by which cells cut and stitch RNA before making proteins. The model predicted the variant would create an abnormal splice site in a brain-specific version of the gene, inserting 13 extra amino acids into the resulting protein (Fortune; DeepMind). A laboratory assay reproduced the predicted splicing change, and the variant was reclassified as likely pathogenic, a formal category that integrates family history, inheritance and functional data (Atlas preprint).
The sequence is instructive. The score ranked the variant. The attribution pointed researchers towards a splicing assay. The assay confirmed the predicted disruption. The model did not independently diagnose the patient, and the published account does not detail all the classification steps (Nature).
How it compares, and what comes next
The Atlas preprint reports retrospective benchmarks against solved rare-disease cases from the GREGoR consortium. AVI placed the known causal variant among a patient's top 50 candidates in 29.5% of cases, versus 12.5% for CADD, the long-established genome-wide scoring tool (Atlas preprint). The advantage was largest in non-coding regions, the vast majority of the genome that sits outside the protein-coding recipes where interpretation has traditionally been hardest. DeepMind researcher Žiga Avsec has acknowledged weaker performance for some enhancer effects, distant DNA stretches that regulate genes from afar (Fortune).
These are ranking comparisons on curated, already-solved cases, produced with DeepMind collaborators and published as a preprint. Finding the known answer retrospectively is a different challenge from increasing diagnoses among genuinely unsolved patients.
On 9 September, Europe's public bioinformatics hub EMBL-EBI announced that the AVI score is now available through the Ensembl Variant Effect Predictor (VEP), the annotation system researchers use to convert raw genetic data into interpretable results (EMBL-EBI). VEP already sits inside the processing pipelines of clinical and research laboratories. Adding AVI means a lab can retrieve the score alongside gene consequences and population frequencies in the same batch operation, without switching to a separate Google portal. VEP makes the score retrievable, without endorsing its clinical accuracy. DeepMind states that AlphaGenome has not been validated or approved for clinical use (DeepMind).
In the reported DNM1 case, AlphaGenome helped researchers prioritise an overlooked variant and supplied a splicing hypothesis that laboratory work could test. Whether that process increases diagnoses at scale depends on prospective studies by independent groups, working with unsolved cases rather than historical answers.
Details about this article
- Model:
- claude-opus-4-6
- Generated:
- 9/10/2026, 2:07:54 AM
- Pipeline run:
- eu_pipeline_20260910_005006
- Watermark:
- SynthID (Google's invisible watermark)
- Human review:
- None before publication